Introduction
Many existing active pharmaceutical ingredients (APIs) (drug compounds) are either weak acids or weak bases; their water solubility can change significantly with the lumen pH changes along the gastrointestinal tract (GIT) as a result of variation in the ionization degree. Severe pH-dependent solubility could pose a great challenge to achieving consistent and predictable performance of oral dosage forms, as abrupt changes in release rate may result in unexpected dissolution, absorption, and bioavailability of the drug, leading to increased risks of adverse side effects or decreased therapeutic efficacy. This problem may be pronounced for weakly basic drugs with low solubility at high pH, especially those with a narrow therapeutic index, or requiring prolonged release, because in the lower GIT the pH is >6.8 [
1]. Furthermore, food intake, disease state (e.g., inflammatory bowel disease, gastritis, colitis), concomitant medication (e.g., proton pump inhibitors), and inter- and intra-individual variations, among other factors, can alter the pH in the GIT, which deviates from the pH of simulated gastric and intestinal fluid for in vitro testing and prediction [
2,
3,
4]. Hence, novel strategies to formulate weakly basic drugs with extreme pH-dependent solubility in controlled release forms could enhance the repertoire of advanced medications available to patients.
To compensate for the varying pH values in the GIT, several approaches have been employed. One approach is the addition of small-molecule pH modifiers to the immediate vicinity of the drug to change the microenvironmental pH (pH
m), thus enhancing the drug solubility. For example, organic acids (e.g., adipic, fumaric, and succinic acids) have been introduced into formulations of weakly basic drugs, reducing the pH
m sufficiently, thereby facilitating drug dissolution, irrespective of the pH of the bulk solvent [
5,
6,
7,
8,
9,
10,
11]. Drug release will depend on the compatibility of the organic acid with the drug in terms of the buffering capacity of organic acids and the pK
a of the drug. Nevertheless, the incorporation of organic acids remains challenging as they are prone to leach out from the formulation, leading to inefficient pH modulation over time [
12]. Therefore, large amounts of organic acids are required in order to achieve prolonged pH-independent drug release [
5,
13], which often makes this approach inadequate for controlled release formulations.
Another approach to compensate for the reduction in drug solubility is the use of enteric coating polymers on tablets, pellets, or beads as permeability modifiers, which can increase drug permeability at higher pH. In such coatings, polymers with pH-dependent solubility or swellability are employed, either alone or incorporated in a hydrophobic polymer (e.g., ethylcellulose (EC)) that acts as a membrane barrier to drug diffusion, resulting in slowed drug release. For example, methacrylic acid–ethyl acrylate copolymer (Eudragit
® L) and hydroxypropyl methylcellulose acetate succinate are frequently incorporated into these membranes as pore forming materials for pH-dependent release [
13,
14,
15,
16,
17,
18,
19,
20,
21,
22,
23]. At a pH above their pKa’s, the pore formers dissolve and leach out of the membrane film to form channels that facilitate drug diffusion. However, as pore formers leach out, the film becomes more porous and weaker, increasing the risk of dose dumping due to weakened structure or ruptures of the coating [
24].
Recent advances using biopolymer-based nanomaterials, molecular imprinted polymers, and mathematical and computational models have been used to address the various challenges of drug delivery systems [
25,
26,
27,
28,
29]. Biopolymers such as starch, collagen, chitosan, etc., are useful for their biocompatibility, biodegradability, and ease of synthesis and modifications [
25]. Controlled release dosage forms capable of being flexible, releasing drugs in a timely manner, with desired duration and dosage, are more important than ever, and mathematical and computational models also function as influential tools in addressing the potential mechanisms or impediments of drug delivery systems [
26,
27,
28,
29].
Previously, a crosslinked terpolymer nanoparticle (TPN), consisting of poly(methacrylic acid)-polysorbate 80-grafted-starch (PMAA-PS 80-g-St) [
30,
31,
32,
33], was incorporated into ethylcellulose films (TPN-EC) to respectively reduce or enhance the permeability of the film coating by a pH-dependent shrinking (at low pH) and swelling (at high pH) mechanism [
34]. Unlike other soluble polymeric pore-formers, the TPN did not increase the viscosity of EC dispersions for coating, attributable to its crosslinking structure [
31,
32,
33]. For the same reason, TPN did not leach out from cast TPN-EC films, maintaining good mechanical properties compared to conventional Eudragit
® L-EC films. When used as a membrane coating over beads loaded with a water-soluble drug, diltiazem HCl, TPN-EC provided faster drug release at pH 6.8 than at pH 1.2 [
34]. The biocompatibility of TPN was demonstrated previously in vitro using isolated rat hepatocytes [
32].
Inspired by previous findings regarding the pH-dependent properties of TPN, in this work, we explore the application of TPN for the first time to develop an advanced, controlled release bilayer-coated bead formulation for weakly basic drugs that exhibits severe pH-dependent water solubility. Verapamil HCl (VER) was selected as a model drug because it undergoes extreme decrease in solubility by several orders of magnitude when the media pH is increased from acidic to neutral and weakly basic conditions [
9,
35]. By exploiting the pH-dependence of TPN, we proposed that an increase in permeability at high pH could help compensate for the low solubility of the drug in its unionized form, thereby permitting a constant release rate when transitioning from gastric to intestinal pH. Additionally, we explored the ability of TPN to serve as a pH
m modifier, owing to the presence of its acidic functional groups in MAA. Because TPN is retained within EC, unlike traditional leachable pH
m modifiers when formulated together with the drug in a matrix, the source of the acidifying agent could be sustained throughout dissolution, while preserving dosage form integrity.
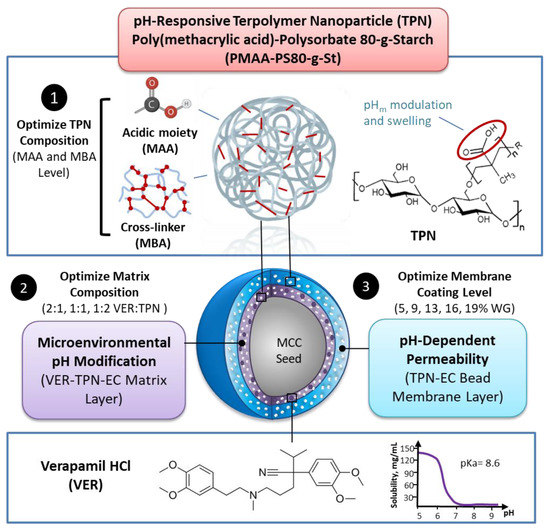
To investigate the effectiveness of combining both the permeability and the pHm modulation strategies of TPN, experiments were performed using EC matrix free-films incorporated with VER and TPN (VER-TPN-EC) and a bilayer-coated bead design consisting of an inner VER-TPN-EC matrix, surrounded by an outer membrane composed of TPN-EC. As illustrated in , the TPN composition was first adjusted by varying the amounts of the pH-responsive monomer, MAA, and cross-linker, N,N′-methylenebis(acrylamide) (MBA) for pH-dependent swelling (medium uptake) and drug release via experiments using composite free-films. The best-performing TPN-containing films were then tested for their ability to lower pHm. Subsequently, the TPN was incorporated into a bilayer-coated bead design, where it was expected to lower the pHm in the matrix layer and regulate permeability in the membrane layer by its pH-dependent swelling. The effects of formulation parameters such as VER:TPN ratio within the matrix layer and membrane coating level on the pH-independence of dissolution rate in various pH conditions were evaluated.
Figure 1. A flow chart of the optimization strategy to achieve pH transition-independent controlled release of VER from a TPN-containing bilayer-coated beads. Formulation strategy for TPN bilayer-coated beads containing weakly basic VER Optimization of TPN composition to achieve pH
m modification and pH-dependent swelling, followed by strategic placement of TPN in the bilayer bead matrix and membrane layers are proposed to overcome pH-dependent solubility of VER. Figure created with
BioRender.com (accessed on 15 December 2022).
Our cumulative research investigating the various applications of TPN (e.g., nanoparticle drug carrier in injectables, enteric coating agent and pore former in film coatings, recrystallization inhibitor in amorphous solid dispersions) is helping us widen the breadth of its capabilities. Namely, in the present research, the ability of TPN for pHm modification, pH-responsive swelling, nanoscale pore formation, interaction with drugs, and non-leachability comprise a set of multi-faceted features that set it apart from traditional excipients, which could improve the efficacy and quality of controlled release dosage forms for weakly basic drugs. As the landscape of new drug molecules continues to shift towards greater challenges (e.g., poor solubility, pH-dependent solubility, narrow therapeutic index, etc.) the need for more advanced, multifunctional excipients can be expected to increase.
 ingredientpharm
 Ingredientpharm